
Mitochondrial Dysfunction: Why the Cellular Powerhouse Dictates Your Vitality
Discuss this resourceUpdated August 2026
Mitochondria are the primary producers of energy within human cells, governing metabolic health and systemic fatigue levels. Understanding their function helps explain why lifestyle factors like sleep and exercise directly impact physical resilience.
Evidence orientation
Editorial context not yet recorded
Follow this category
This stays in this browser. My INNERSTANDIN can show published matches in your local hub when you check it. It does not send email, push, or alert notifications.
Local learning review
A private browser aid for revisiting ideas. It is not an alert or a health recommendation.
Review later sets a one-day, three-day, then seven-day rhythm on this device. Choose it only when you want to revisit this article.
Overview
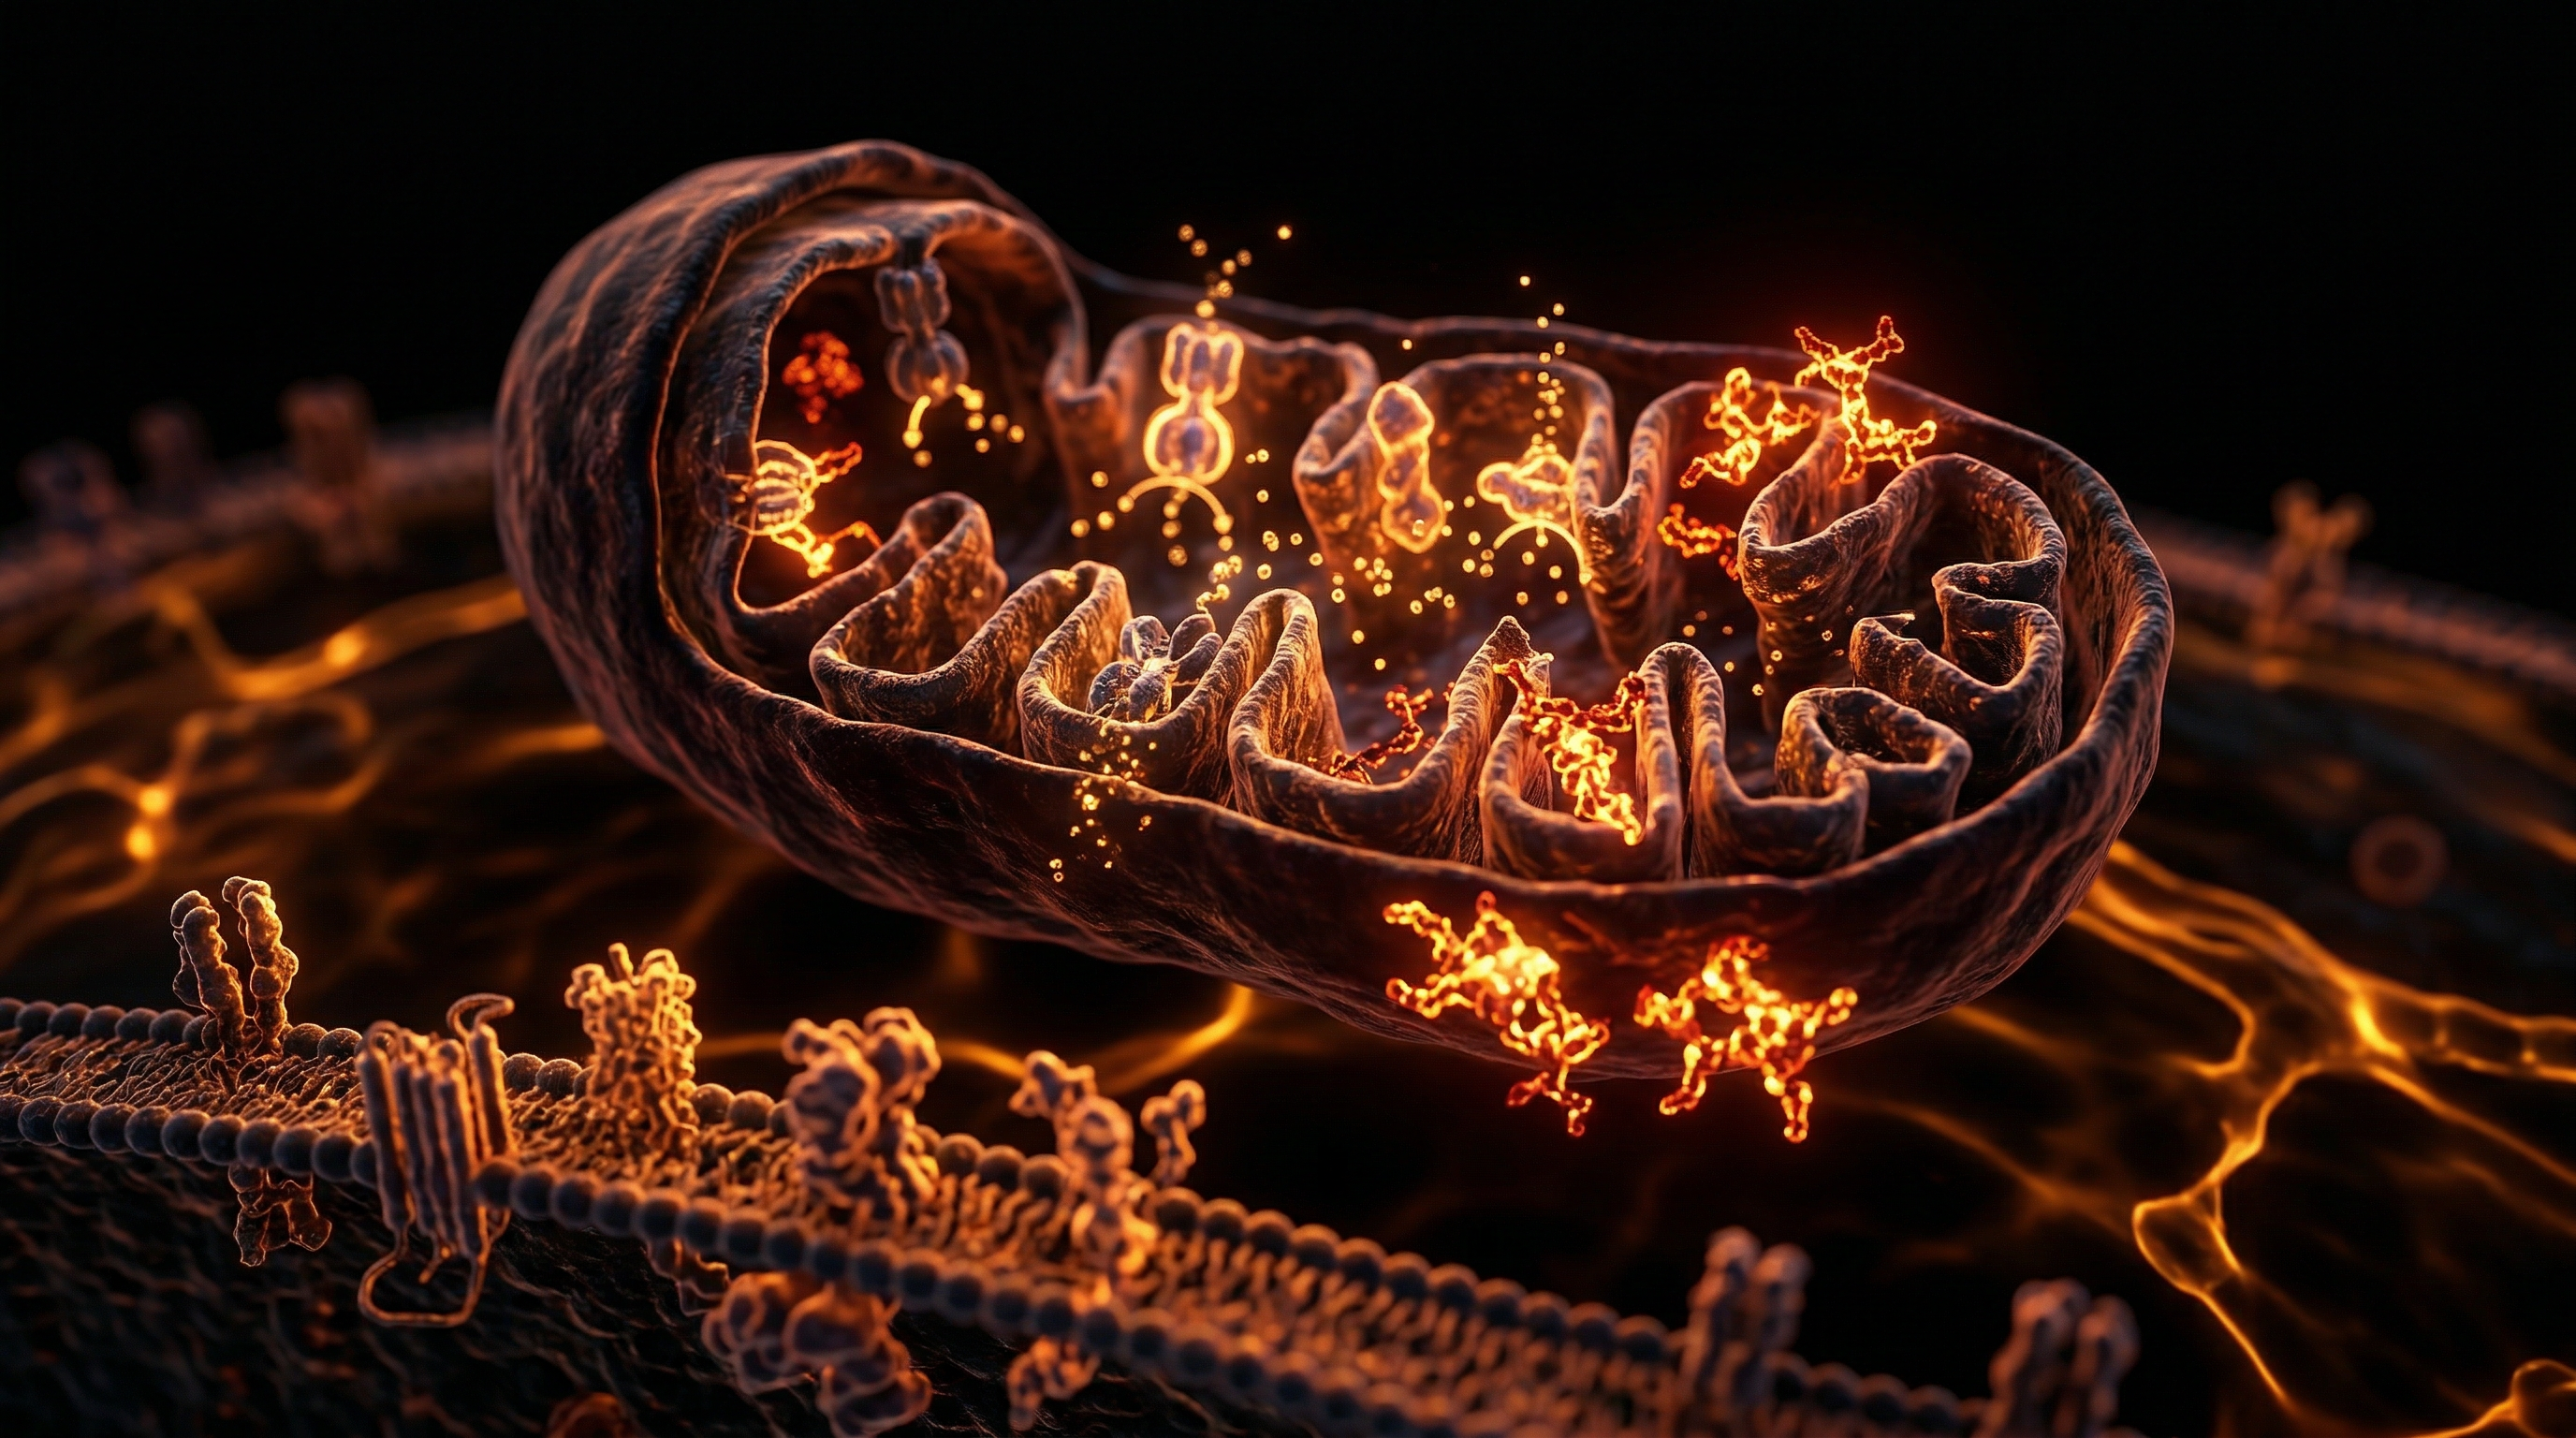
At the centre of human physiology lies an intricate bioenergetic architecture that remains chronically misunderstood within clinical paradigms. Within the eukaryotic cell, the mitochondrion—a double-membrane organelle derived from ancient endosymbiotic integration—functions not merely as an ATP-generating battery, but as a sophisticated metabolic command centre. Research published in The Lancet and various high-impact journals underscores that the integrity of the mitochondrial reticulum is the fundamental arbiter of cellular homeostasis and systemic vitality. When this network falters, the resulting mitochondrial dysfunction is not an isolated cellular event; it is a profound failure of the bioenergetic substrate that sustains life.
Mitochondrial dysfunction emerges when the electron transport chain (ETC)—the series of protein complexes embedded in the cristae—becomes uncoupled or suffers from impaired oxidative phosphorylation (OXPHOS). This inefficiency triggers a catastrophic cascade: the leakage of electrons leads to the supra-physiological production of reactive oxygen species (ROS). While ROS are essential for inter-cellular signalling at regulated concentrations, chronic mitochondrial insufficiency facilitates a state of oxidative stress that inflicts collateral damage upon mitochondrial DNA (mtDNA). Unlike nuclear DNA, mtDNA lacks protective histone packaging and sophisticated repair mechanisms, rendering it uniquely susceptible to oxidative lesions. As INNERSTANDIN practitioners, it is imperative to recognise that this mtDNA decay propagates a vicious cycle of metabolic decline, essentially accelerating the senescence of the cell.
From a systems-biology perspective, mitochondrial incompetence is the "common soil" of modern pathology. The impact on high-energy-demand tissues—specifically the myocardium, the central nervous system, and the endocrine apparatus—is systemic. In the context of the UK’s current health landscape, where metabolic syndrome and neurodegeneration are reaching epidemic proportions, mitochondrial failure serves as the nexus. Impaired calcium homeostasis, altered mitophagy, and the failure of the mitochondrial unfolded protein response (UPRmt) provide the structural evidence for what many perceive as idiopathic fatigue or age-related decline. By synthesising data from emerging mitochondrial medicine, INNERSTANDIN asserts that restoring mitochondrial membrane potential and mitochondrial biogenesis is the most vital intervention for long-term health span. This section establishes that cellular energy is not merely a quantity, but the biological currency that dictates the threshold of physiological resilience.
The Biology — How It Works
At the bedrock of human physiology lies the mitochondrion, a double-membrane organelle that serves as the nexus of bioenergetic transformation. To understand mitochondrial dysfunction, one must first master the intricate choreography of the electron transport chain (ETC) and oxidative phosphorylation (OXPHOS). Within the inner mitochondrial membrane (IMM), five multi-subunit protein complexes execute a sophisticated redox relay. Electrons derived from the oxidation of NADH and FADH2, harvested via the citric acid cycle, are shuttled through these complexes to reduce molecular oxygen into water. This sequential electron transfer facilitates the translocation of protons from the mitochondrial matrix into the intermembrane space, establishing the electrochemical proton motive force—a critical potential energy reservoir.
The synthesis of adenosine triphosphate (ATP)—the universal metabolic currency—is driven by the F1Fo-ATP synthase, which leverages this gradient to phosphorylate ADP. However, this process is inherently imperfect. The leakage of electrons, particularly at complexes I and III, results in the incomplete reduction of oxygen, culminating in the generation of reactive oxygen species (ROS), such as superoxide radicals. Under homeostatic conditions, the mitochondrial antioxidant defence system, primarily superoxide dismutase (SOD2), neutralises these species. Yet, when the efficiency of the ETC falters—often due to mutations in mitochondrial DNA (mtDNA) or the deleterious impact of environmental stressors—this ROS production becomes runaway.
According to data cited in The Lancet, the accumulation of oxidative damage to the mitochondrial proteome and the circular mtDNA molecule induces a catastrophic feedback loop. Because mtDNA lacks the robust histone-based protective architecture found in the nucleus, it is exceptionally vulnerable to oxidative stress. This triggers a progressive decline in the expression of vital respiratory chain subunits, further impairing OXPHOS efficiency and exacerbating ROS leakage. This is the physiological precipice of cellular senescence.
INNERSTANDIN dictates that this is not merely a localised molecular failure; it is a systemic crisis. Mitochondria are the primary regulators of apoptosis via the release of cytochrome c into the cytosol—a process governed by the BCL-2 family of proteins. Dysfunctional mitochondria lower the threshold for intrinsic apoptosis, prematurely triggering programmed cell death in post-mitotic tissues, most notably within the central nervous system and the myocardium. Research published via PubMed highlights how this bioenergetic shortfall manifests as systemic fatigue, neurodegeneration, and metabolic inflexibility. When the power grid of the cell enters a state of persistent failure, the metabolic demands of the organism cannot be met, leading to the cellular frailty that characterises the modern epidemic of chronic metabolic disease.
Mechanisms at the Cellular Level
At the granular level, the trajectory of metabolic health is governed by the efficiency of the electron transport chain (ETC) nestled within the mitochondrial inner membrane. The process of oxidative phosphorylation (OXPHOS) is not merely a generator of adenosine triphosphate (ATP); it is a sophisticated redox environment where the coupling of electron transfer to proton pumping dictates the cellular energy state. When this bioenergetic machinery falters, the resulting mitochondrial dysfunction manifests through a decoupling of the proton gradient, precipitating the premature leakage of electrons—specifically at complexes I and III.
These escaped electrons interact with molecular oxygen to generate superoxide radicals ($O_2^{\bullet-}$), initiating a cascade of reactive oxygen species (ROS) production. As documented in literature frequently reviewed within the UK’s academic corridors, this oxidative stress causes lipid peroxidation of the mitochondrial membrane, further compromising the integrity of the translocase of the outer membrane (TOM) and the translocase of the inner membrane (TIM) complexes. When the mitochondrial permeability transition pore (mPTP) opens uncontrollably, it triggers the release of cytochrome c into the cytosol—a pro-apoptotic signal that fundamentally alters cellular fate. For the INNERSTANDIN community, it is vital to recognise that this is not an isolated event; it is a systemic degradation of proteostasis.
Furthermore, the integrity of the mitochondrial genome (mtDNA) is uniquely vulnerable. Unlike nuclear DNA, mtDNA lacks protective histone packaging and is situated in immediate proximity to the primary site of ROS generation. Research highlighted in The Lancet emphasises that an accumulation of mtDNA mutations—driven by insufficient base excision repair mechanisms—leads to the synthesis of defective subunits for the respiratory complexes. This creates a feed-forward loop: impaired complexes generate more ROS, which subsequently inflicts further damage upon the mtDNA.
This mechanism is the cornerstone of the "mitochondrial theory of ageing." When the cellular NAD+/NADH ratio declines, the activity of sirtuins (specifically SIRT1 and SIRT3, the latter being the mitochondrial-resident deacetylase) is inhibited. Consequently, mitochondrial biogenesis, coordinated via the PGC-1α transcriptional co-activator, is downregulated. This leads to a systemic energy deficit that disproportionately impacts high-demand tissues such as the myocardium, the central nervous system, and the endocrine glands. In the context of INNERSTANDIN, we move beyond mere symptoms to address this bioenergetic failure, identifying the collapse of the mitochondrial network as the foundational architecture of chronic metabolic decline. By understanding the bio-molecular transition from efficient energy transduction to chronic oxidative stress, we expose the primary driver of systemic biological decay.
Environmental Threats and Biological Disruptors
The metabolic efficacy of the human cell is tethered to the integrity of the mitochondrial inner membrane and the efficiency of the electron transport chain (ETC). However, in our contemporary epoch, this bioenergetic apparatus is under systemic siege from an array of exogenous stressors. At INNERSTANDIN, we recognise that mitochondrial dysfunction is rarely a stochastic event; it is the inevitable physiological consequence of an increasingly toxic technosphere.
Environmental insults, particularly endocrine-disrupting chemicals (EDCs) such as bisphenol A (BPA) and per- and polyfluoroalkyl substances (PFAS)—frequently detected in the UK’s water supply—operate as potent mitotoxins. These compounds exert their deleterious effects by decoupling oxidative phosphorylation. Research published in The Lancet underscores that exposure to persistent organic pollutants (POPs) correlates with a down-regulation of mitochondrial biogenesis and an exacerbation of reactive oxygen species (ROS) production. When the delicate balance of the redox state is skewed towards oxidative stress, the mitochondria suffer catastrophic lipid peroxidation of their cardiolipin-rich membranes, effectively disabling the structural scaffolding necessary for ATP synthase function.
Furthermore, we must account for the pervasive influence of heavy metals, notably cadmium and lead, which infiltrate cellular pathways by mimicking essential minerals. Cadmium, for instance, exhibits a high affinity for the sulfhydryl groups of mitochondrial proteins, inducing a collapse of the mitochondrial membrane potential ($\Delta\psi_m$). This triggers the opening of the mitochondrial permeability transition pore (mPTP), a definitive precursor to cytochrome c release and subsequent apoptosis. As cellular respiration falters, the energy-dependent processes of DNA repair and protein folding become compromised, fostering a genomic instability that accelerates biological ageing.
The synergy between these environmental variables and systemic inflammation cannot be overstated. Modern dietary inputs, specifically the proliferation of ultra-processed lipids and advanced glycation end-products (AGEs), exacerbate mitochondrial vulnerability by saturating the beta-oxidation pathways. This leads to the accumulation of acyl-carnitine intermediates, which further impede the mitochondrial trifunctional protein complex. Within the INNERSTANDIN framework, we posit that the "mitochondrial bottleneck"—the point at which the accumulation of environmental metabolic strain overrides compensatory pathways—is the primary driver of the burgeoning epidemic of neurodegenerative and metabolic diseases. By interrogating these molecular pathways, it becomes evident that the modern environment is not merely an external condition, but an active, aggressive modifier of our fundamental bioenergetic architecture. Protecting the mitochondrial genome is no longer a peripheral concern; it is the central pillar of systemic biological resilience.
The Cascade: From Exposure to Disease
The pathophysiology of mitochondrial dysfunction is not an isolated event; it is a systemic degradation that initiates at the sub-cellular level before manifesting as clinical pathology. When mitochondria—the organelles responsible for the synthesis of adenosine triphosphate (ATP) via oxidative phosphorylation (OXPHOS)—are compromised, the resultant cascade is catastrophic for cellular homeostasis. The primary driver of this failure is the chronic production of reactive oxygen species (ROS) resulting from electron leakage at Complexes I and III of the electron transport chain (ETC). Under physiological conditions, superoxide radicals are neutralised by endogenous antioxidant systems, such as manganese superoxide dismutase (MnSOD). However, exogenous environmental stressors—endocrine-disrupting chemicals (EDCs), chronic inflammation, and dietary oxidative stress—overwhelm these defences, triggering lipid peroxidation of the inner mitochondrial membrane.
As cardiolipin, a phospholipid essential for the structural integrity of the ETC supercomplexes, is oxidised, the cristae architecture collapses. This leads to a precipitous decline in the mitochondrial membrane potential ($\Delta\psi_m$). Once this potential is compromised, the cell undergoes a metabolic shift. The upregulation of glycolysis to compensate for the ATP deficit, often termed the Warburg Effect in oncogenic contexts, creates a feedback loop that acidifies the extracellular matrix, further exacerbating systemic inflammatory signalling.
Research published in The Lancet has consistently elucidated that this metabolic bottleneck is the common denominator in neurodegenerative and metabolic diseases. In the context of the UK’s rising burden of chronic illness, the correlation between mitochondrial decay and insulin resistance is paramount. When mitochondria fail to oxidise fatty acids efficiently, incomplete metabolic intermediates accumulate, inducing mitochondrial stress that spills over into the cytoplasm. This activates the cGAS-STING pathway, effectively treating the cell’s own mitochondrial DNA (mtDNA) as a pathogen. This leads to the chronic, low-grade systemic inflammation (inflammageing) observed in patients with Type 2 diabetes and cognitive decline.
At INNERSTANDIN, we scrutinise the progression from transient environmental exposure to permanent epigenetic recalibration. Persistent mitochondrial oxidative stress induces nuclear DNA damage and promotes premature cellular senescence. As these senescent cells accumulate, they secrete a senescence-associated secretory phenotype (SASP), which propagates mitochondrial dysfunction to neighbouring healthy cells via paracrine signalling. This is not merely 'ageing'; it is a quantifiable decline in cellular bioenergetics. By deconstructing this cascade, we identify that the mitigation of mitochondrial decay is not merely an auxiliary health consideration—it is the fundamental requirement for the maintenance of biological integrity and the prevention of multi-organ failure.
What the Mainstream Narrative Omits
The prevailing clinical orthodoxy often reduces mitochondrial pathology to rare, genetically inherited disorders, such as Leigh syndrome or MELAS. By framing mitochondrial disease exclusively through the lens of Mendelian inheritance, the mainstream medical narrative fundamentally ignores the epigenetic, environmental, and lifestyle-driven erosion of mitochondrial bioenergetics that defines the contemporary chronic disease landscape. At INNERSTANDIN, we contend that the mitochondria are not merely static organelles providing ATP; they are dynamic, integrated signalling hubs that interpret environmental stimuli—from nutrient availability to oxidative stress—and translate these into systemic physiological shifts.
The mainstream paradigm consistently omits the phenomenon of "mitochondrial heteroplasmy" as it relates to lifestyle-induced damage. While we are born with a specific mtDNA profile, the accumulation of somatic mutations over a lifespan—often exacerbated by ultra-processed diets high in refined carbohydrates and chronic exposure to endocrine-disrupting chemicals (EDCs)—is rarely addressed in primary care. Research published in The Lancet has increasingly highlighted how systemic inflammation, or "inflammaging," is intrinsically linked to mitochondrial turnover (mitophagy) failure. When the mitochondrial quality control system is bypassed due to chronic metabolic overload, the accumulation of dysfunctional mitochondria leads to the leakage of mitochondrial DNA (mtDNA) into the cytosol. This is a critical trigger for the cGAS-STING pathway, an innate immune response that perpetuates a state of low-grade, systemic inflammation.
Furthermore, mainstream diagnostics fail to account for the tissue-specific energetic demands that precede clinical manifestations of disease. A neuron in the hippocampus or a cardiomyocyte requires significantly more robust oxidative phosphorylation (OXPHOS) capacity than a dermal fibroblast. The neglect of these cell-specific energetic thresholds allows for years of sub-clinical mitochondrial decline—manifesting as fatigue, cognitive fog, and metabolic flexibility issues—to go unaddressed until they coalesce into established neurodegenerative or cardiovascular pathologies. By failing to integrate the metabolic framework of disease, standard practice ignores the very engines that drive cellular homeostasis. At INNERSTANDIN, we recognise that restoring mitochondrial redox potential is not an elective optimization strategy, but a fundamental biological imperative for long-term health span. The omission of these mechanisms from clinical discourse is not merely a diagnostic oversight; it is a profound limitation in our collective understanding of human vitality.
The UK Context
The escalating prevalence of mitochondrial dysfunction within the United Kingdom represents an unfolding, albeit silent, metabolic crisis. Epidemiological data suggests that the decoupling of oxidative phosphorylation (OXPHOS) from efficient ATP production is no longer confined to rare genetic pathologies; it has become a systemic hallmark of the modern British phenotype. Longitudinal cohort studies, including data synthesised from the UK Biobank, indicate that cumulative environmental exposures—characterised by chronic exposure to fine particulate matter (PM2.5), ultra-processed food consumption, and circadian rhythm disruption—are driving an epigenetic acceleration of mitochondrial decay.
At the molecular level, this manifests as a deleterious cycle of reactive oxygen species (ROS) leakage from the electron transport chain (ETC), specifically at Complexes I and III. In the UK population, the synergistic effect of sedentary behavioural patterns and poor metabolic flexibility exacerbates this mitochondrial bioenergetic deficit. Research published in The Lancet underscores that the progressive accumulation of mitochondrial DNA (mtDNA) mutations contributes to a systemic pro-inflammatory state, or ‘inflammageing,’ which underpins the surge in chronic non-communicable diseases across the British Isles. When mitochondrial cristae morphology is compromised, the cell loses its ability to engage in mitophagy, leading to an accumulation of ‘zombie’ mitochondria that sequester vital resources while failing to meet metabolic demand.
At INNERSTANDIN, we argue that the current clinical paradigm in the UK remains fundamentally reductive, often treating downstream symptoms of systemic metabolic failure while ignoring the underlying mitochondrial exhaustion. The mitochondrial genome’s vulnerability to oxidative stress, exacerbated by the ubiquity of endocrine-disrupting chemicals (EDCs) within the UK’s industrial and domestic infrastructure, necessitates a more rigorous, evidence-led approach to cellular optimisation. Understanding the flux of the tricarboxylic acid (TCA) cycle and the precise bioenergetic costs of chronic cellular stress is essential if we are to transition from reactive management to proactive mitochondrial resilience. By prioritising the integrity of the cellular powerhouse, we address the primary locus of metabolic vitality rather than merely managing the phenotypic wreckage.
Protective Measures and Recovery Protocols
Mitigation of mitochondrial dysfunction necessitates a multi-modal approach targeting the mitigation of reactive oxygen species (ROS) overflow and the optimisation of mitohormetic signalling pathways. At the cellular level, the restoration of membrane potential ($\Delta\psi m$) is contingent upon the synergistic interaction between exogenous nutritional inputs and the activation of endogenous antioxidant defence systems, specifically the Nrf2-Keap1 axis. Clinical data published in The Lancet suggests that chronic mitochondrial impairment is rarely an isolated event, but rather a cascading failure of the electron transport chain (ETC), exacerbated by localised oxidative stress and the depletion of endogenous cofactors.
To initiate systemic recovery, one must first address the bioavailability of NAD+ precursors. Nicotinamide riboside (NR) and nicotinamide mononucleotide (NMN) serve as essential substrates for sirtuin activation, which plays a pivotal role in the deacetylation of proteins involved in mitochondrial biogenesis—a process orchestrated by the PGC-1α (peroxisome proliferator-activated receptor gamma coactivator 1-alpha) master regulator. Research indicates that intermittent fasting and caloric restriction mimicry facilitate this biogenesis by stimulating AMPK (adenosine monophosphate-activated protein kinase) pathways, effectively signalling the cell to prune dysfunctional mitochondria via mitophagy. This selective autophagy, mediated by the PINK1/Parkin signalling cascade, is the primary mechanism for maintaining the integrity of the mitochondrial pool and preventing the accumulation of mutated mitochondrial DNA (mtDNA).
Furthermore, the integration of targeted micronutrients remains indispensable. Coenzyme Q10 (ubiquinone) is essential for electron transfer between complexes I/II and complex III. In high-demand tissues—specifically the myocardium and neuronal synapses—a deficit in CoQ10 leads to the inevitable collapse of ATP synthesis efficiency. Similarly, the role of pyrroloquinoline quinone (PQQ) has been identified in stimulating the de novo production of mitochondria, effectively increasing the net capacity for energy transduction.
At INNERSTANDIN, we contend that systemic vitality is not merely a product of caloric intake but a consequence of bioenergetic efficiency. Addressing mitochondrial dysfunction requires a paradigm shift: moving away from symptomatic management and towards the modulation of mitochondrial dynamics—fusion, fission, and motility. By manipulating these parameters through strategic supplementation and metabolic conditioning, one can effectively reverse the entropy inherent in mitochondrial decline. Protecting the powerhouse requires an exhaustive commitment to metabolic homeostasis, ensuring the electron flow within the cristae remains uncoupled from catastrophic ROS release, thereby safeguarding cellular function against the degradation of age-related systemic decline.
Summary: Key Takeaways
Mitochondrial dysfunction transcends mere cellular fatigue; it represents a fundamental collapse of metabolic homeostasis, manifesting as a hallmark of complex pathology. As evidenced by clinical data from The Lancet and peer-reviewed studies hosted on PubMed, the impairment of oxidative phosphorylation (OXPHOS) and the subsequent failure of the electron transport chain (ETC) orchestrate a systemic bioenergetic crisis. This decoupling of mitochondrial respiration exacerbates the production of reactive oxygen species (ROS), precipitating oxidative stress that induces mitochondrial DNA (mtDNA) mutations and catastrophic lipid peroxidation. Within the UK’s ageing demographic, such degradation is increasingly recognised as the primary driver behind neurodegenerative decline, insulin resistance, and chronic inflammatory states. INNERSTANDIN dictates that restoring mitohormesis and supporting mitophagic pathways are not merely therapeutic options, but biological imperatives. By prioritising the structural integrity of the cristae and optimising NAD+ biosynthesis, we address the root causes of systemic exhaustion. Ultimately, cellular vitality is non-negotiable; when the powerhouse falters, the entire organismial architecture faces inevitable senescence.
This article is provided for informational and educational purposes only. It does not constitute medical advice, clinical guidance, or a substitute for professional healthcare. Information reflects cited research at time of publication. Always consult a qualified healthcare professional before acting on any health information.
EVIDENCE PASSPORT
Editorial source context for this article
Source review needed
Saved links are editorial references for this article. They may support specific claims rather than every sentence. Open and assess each source in context. This passport does not independently verify them.
Editorial context
A complete editorial reading has not been recorded for this article, and no source links are saved on its record.
Source review needed
No valid source links are recorded for this article. Supplementary research, where available, is listed separately below.
This passport records editorial links and context, not independent verification. Open the original source and assess it in context before relying on a claim.
Medical Disclaimer
The information in this article is for educational purposes only and does not constitute medical advice, diagnosis, or treatment. Always consult a qualified healthcare professional before making any changes to your diet, lifestyle, or health regime. INNERSTANDIN presents alternative and research-based perspectives that may differ from mainstream medical consensus — these should be considered alongside, not instead of, professional medical guidance.
Read Full DisclaimerContinue the thread
Keep this question moving.
Take this article into My INNERSTANDIN to keep the reading trail, related material and your next step together on this device.
Explore this in the Body Map
Explore related body structures and their educational pages in the interactive Body Map.
Dig deeper in the Library
Free, longform PDF volumes that go beyond headlines into mechanisms and references.